
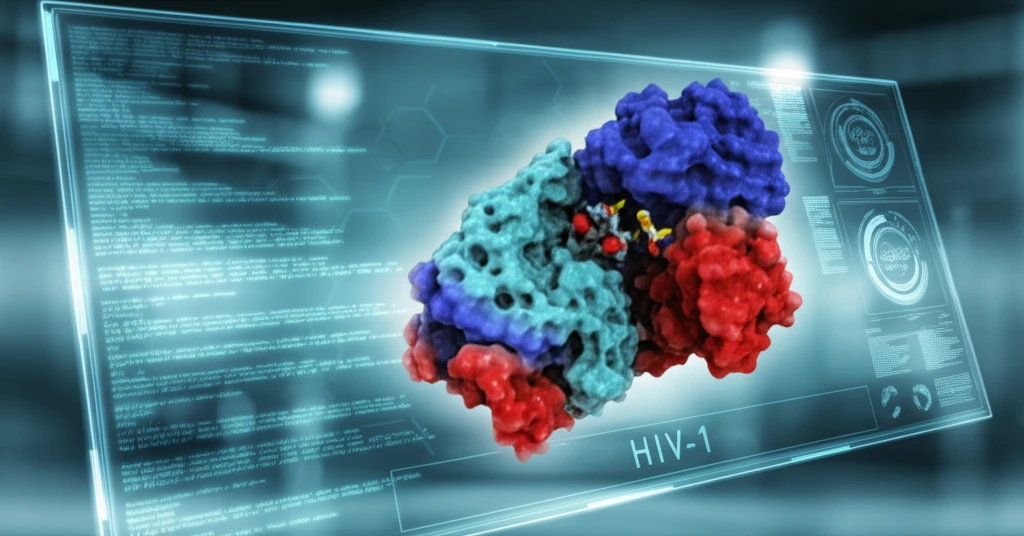
Unlocking the Secrets: How Structure-Based Virtual Screening is Revolutionizing HIV-1 Integrase Inhibitor Design

"A combined approach using virtual screening and molecular dynamics simulation offers a powerful strategy for identifying potent HIV-1 integrase inhibitors."
The relentless pursuit of effective treatments for Human Immunodeficiency Virus type 1 (HIV-1) continues to drive innovation in drug discovery. HIV-1, the pathogen responsible for Acquired Immunodeficiency Syndrome (AIDS), necessitates continuous therapeutic intervention to manage its devastating effects on the human immune system. As one of the world’s major epidemics, particularly affecting populations in Africa, finding new ways to combat HIV-1 remains a critical global health priority.
A key enzyme in the HIV-1 lifecycle is integrase (IN), which is essential for the virus to integrate its genetic material into the host cell's DNA. This integration process is a crucial step for HIV-1 to replicate and establish a persistent infection. Consequently, HIV-1 integrase has become a prime target for antiviral drug development. Inhibiting integrase can disrupt the viral replication cycle, offering a pathway to control the infection.
Traditional drug discovery methods are often time-consuming and resource-intensive. However, advancements in computational techniques have opened new avenues for accelerated drug design. In silico approaches, such as structure-based virtual screening and molecular dynamics simulations, are increasingly being used to identify potential drug candidates with greater efficiency and precision. These methods allow researchers to explore vast chemical spaces and predict the binding affinity of molecules to target proteins like HIV-1 integrase.
HIV’s Ongoing Health Impact
HIV attacks the body's immune system and, without treatment, can lead to AIDS. The CDC states that there is currently no effective cure and that people who acquire HIV have it for life, although proper medical care can control the virus. HIV.gov similarly explains that untreated HIV can make it harder for the body to fight infections and diseases and may progress to AIDS when the immune system is badly damaged. These realities sustain the need for research into improved prevention and treatment strategies.
Treatment and Its Limits
HIV infection is managed through medical care intended to control the virus rather than eliminate it. Mayo Clinic reports that HIV makes the body less able to clear infections and diseases, and that untreated infection can eventually cause AIDS. The source also notes that HIV spreads through contact with some body fluids. These limitations help explain why continued research into new antiviral targets and drug-design strategies remains important.
A Continuing Research History
The history of HIV research can be understood as an ongoing effort to prevent, treat, and ultimately better control infection. Foundational work has included studying HIV as an infectious agent and understanding AIDS as the disease caused by the virus. The specific milestones relevant to integrase inhibitor design require evidence from dedicated historical sources, which are not provided here.
Structure-Based Virtual Screening: A Powerful Tool for HIV-1 Integrase Inhibitor Discovery
Structure-based virtual screening is a computational technique that uses the three-dimensional structure of a target protein, such as HIV-1 integrase, to identify molecules that are likely to bind to it. This approach involves screening large libraries of chemical compounds and predicting their binding affinity based on their structural complementarity to the target protein's active site. By prioritizing compounds with high binding affinity, researchers can significantly reduce the number of compounds that need to be synthesized and tested experimentally.
- Target Preparation: Obtaining or generating a high-resolution structure of the target protein, ensuring it is properly prepared for docking simulations.
- Database Selection: Choosing a suitable database of chemical compounds to screen. These databases can contain millions of molecules with diverse structures and properties.
- Docking Simulations: Using computational algorithms to predict the binding mode and affinity of each compound in the database to the target protein.
- Scoring and Ranking: Evaluating the docking results using scoring functions to rank the compounds based on their predicted binding affinity.
- Visual Inspection: Manually inspecting the top-ranked compounds to assess their binding interactions and identify potential lead candidates.
The Research Landscape
Current HIV research broadly encompasses efforts to prevent, treat, or cure HIV/AIDS, as well as fundamental studies of the virus and the disease it causes. Structure-based virtual screening represents one possible research direction within that larger effort. Without subsection-specific sources, its latest findings, study results, and review conclusions cannot be stated reliably.
Limits and Uncertainties
Research approaches for HIV-1 integrase inhibitor design may encounter scientific and practical limitations, but the available material does not identify particular failures or counterarguments. It therefore would be premature to attribute specific shortcomings to structure-based virtual screening. A balanced assessment would require sources reporting unsuccessful designs, reproducibility concerns, resistance issues, or clinical limitations.
Comparing Design Strategies
Structure-based virtual screening can be compared with other HIV-1 integrase inhibitor discovery methods only when evidence describes their procedures and outcomes on a common basis. No comparison-specific sources are provided here. Accordingly, claims about relative speed, accuracy, cost, or clinical performance would require additional supporting research.
The Future of HIV-1 Integrase Inhibitor Design
The convergence of computational power and structural biology is transforming the landscape of drug discovery. Structure-based virtual screening, augmented by molecular dynamics simulation, offers a powerful and efficient strategy for identifying potential HIV-1 integrase inhibitors. While further experimental validation is essential, these in silico techniques hold immense promise for accelerating the development of novel antiviral therapies to combat HIV-1 infection.
Interpreting the Opportunity
The supplied material establishes the continuing importance of HIV research and the need for medical care that controls, rather than cures, infection. It does not provide expert commentary specifically evaluating structure-based virtual screening for HIV-1 integrase inhibitors. Any synthesis about the method’s significance should therefore remain provisional until supported by dedicated expert or review sources.
Future Research Directions
Future work in HIV-1 integrase inhibitor design may investigate improved ways to discover and optimize compounds through structural information. However, the available subsection material does not identify particular emerging targets, technologies, clinical candidates, or timelines. Specific predictions about the field’s next frontiers would require sources focused on those developments.
Beyond Molecular Design
HIV research exists within a broader health context in which infection can damage immune function and untreated disease can progress to AIDS. The available material does not address systemic challenges such as research access, healthcare delivery, affordability, or global disparities. Those issues should not be characterized in detail without sources specifically examining them.
Patient-Relevant Stakes
The human stakes of HIV research are reflected in the fact that HIV can persist for life and, without treatment, can lead to AIDS. Effective medical care can control the virus, making continued treatment development relevant to people living with HIV. The supplied material does not provide patient narratives or evidence about how structure-based screening specifically affects real-world outcomes.