
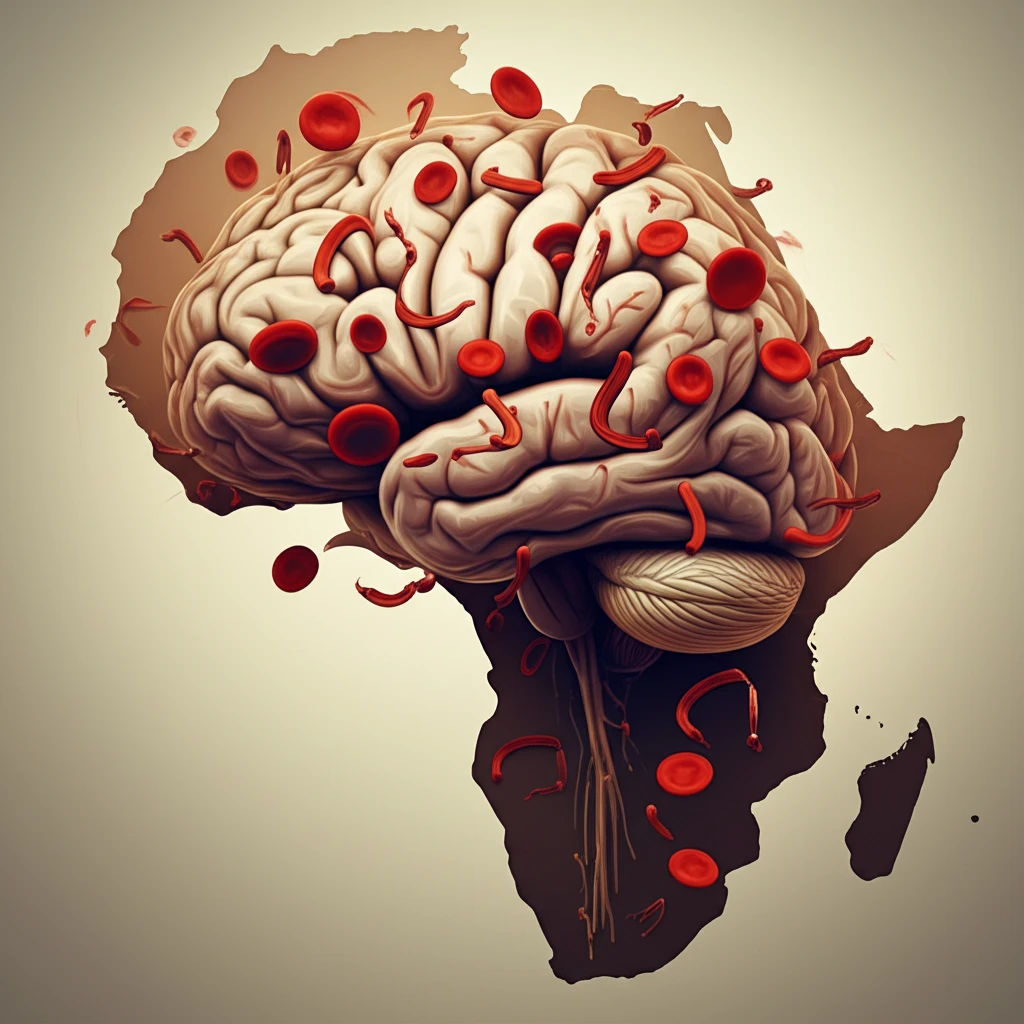
Sickle Cell Disease and the Brain: Unmasking Neurological Risks in Africa
"A systematic review reveals the silent threats of sickle cell disease, calling for proactive screening and management strategies to protect vulnerable populations."
Sickle cell disease (SCD) is a significant public health concern in Africa, where it's associated with high rates of illness and death. Beyond the well-known physical symptoms, SCD can lead to serious neurological complications, impacting patients' quality of life and placing a heavy burden on families. Unfortunately, there are gaps in our understanding of how SCD affects the brain in African populations.
Imagine a child with SCD struggling not only with pain crises but also with silent strokes or cognitive difficulties that go unnoticed. These neurological issues can lead to learning disabilities, behavioral problems, and reduced overall well-being. Because these complications aren't always obvious, they often go unmanaged, leading to long-term consequences.
To address this critical gap, researchers conducted a systematic review, pooling data from various studies to estimate the prevalence of neurological complications in African individuals with SCD. The goal is to provide a clearer picture of the problem, inform healthcare policies, and guide future research efforts to improve the lives of those affected.
Neurological Burden
The CDC provides data on sickle cell disease prevalence, complications, and related health impacts. Research using the St George's Hospital Red Cell Database has specifically examined neurological complications in adults with sickle cell disease. A systematic review of central nervous system complications included studies reporting outcomes for at least 15 individuals, while a 2024 analysis found no statistically significant difference in CNS complication likelihood among most sickle cell disorder types, although seizures were less likely in SC and ST disease and multiple comorbidities were associated with worse outcomes.
Established Care
Clinical care for neurological complications of sickle cell disease generally relies on established diagnostic, preventive, and treatment practices. However, the available material does not identify a single standard approach or provide enough evidence to compare its limitations. Any assessment of these methods should therefore account for differences in patient populations, clinical resources, and local implementation.
Foundational Understanding
The neurological risks of sickle cell disease have become an increasingly recognized part of the condition's broader clinical history. Foundational progress appears to have developed through improved recognition of brain complications and the gradual refinement of clinical responses. The supplied material does not provide specific dates, discoveries, or milestone studies, so a detailed historical timeline cannot be established here.
The Silent Threat: Unveiling Neurological Complications
Neurological complications in SCD arise from vaso-occlusion and hemolysis. The polymerization of deoxy-hemoglobin S results in inflexible, sickle-shaped red blood cells that block small blood vessels. This can cause a range of issues:
- Stroke (Ischemic or Hemorrhagic): Blocked blood flow deprives the brain of oxygen, causing damage.
- Silent Cerebral Infarcts: Small, often unnoticed areas of damage that can accumulate over time.
- Cognitive Impairment: Difficulties with memory, attention, and learning.
- Seizures: Disrupted electrical activity in the brain.
- Chronic Headaches: Persistent pain that can affect concentration and quality of life.
Research Landscape
Current scholarship includes reviews and observational analyses focused on the broad neurological spectrum of sickle cell disease. Recent work has examined adult neurological complications, the burden of CNS complications, and whether complication risks differ among sickle cell disorder types. The supplied material does not identify a single definitive new intervention or consensus beyond these research directions.
Limits and Variation
Evidence does not support assuming that every sickle cell disorder carries identical neurological risks: a 2024 analysis found no statistically significant difference in overall CNS complication likelihood across most types, but reported lower seizure odds in SC and ST disease. At the same time, multiple comorbidities were associated with worse outcomes, showing why genotype alone may not explain neurological prognosis. Reviews emphasize that CNS involvement is associated with substantial morbidity and mortality, while also documenting improved diagnostic and treatment options and major advances in investigation and prevention.
Comparing Risk
Comparisons of neurological risk in sickle cell disease should distinguish among disease types, complications, comorbidities, and patient populations. The supplied material suggests that broad differences between sickle cell disorder types may be limited, while particular outcomes such as seizures can still vary. More specific comparisons require consistent definitions, sufficiently large cohorts, and comparable follow-up.
Turning Knowledge into Action: A Call for Proactive Care
This systematic review highlights the urgent need for increased awareness and proactive management of neurological complications in African individuals with SCD. By quantifying the burden of these complications, we can inform policymakers and healthcare providers, leading to better screening programs and treatment strategies.
Integrated Perspective
Taken together, the material presents sickle cell disease as a systemic condition with a clinically important neurological dimension. Evidence spans adult database studies, systematic reviews, genotype comparisons, and reviews of diagnostic and treatment advances. A balanced interpretation should recognize both meaningful progress and continuing uncertainty about how risks vary across patients and settings.
Beyond Overt Stroke
Future work must look beyond overt stroke, because cerebral complications may remain unnoticed despite a high prevalence and substantial effects on quality of life. Protecting brain function is a central priority because sickle cell disease can produce multiple cerebrovascular complications that threaten neurocognitive function and life. These priorities support research aimed at detecting less visible cerebral injury and improving strategies that preserve neurological function.
Systemic Neurological Risk
People with sickle cell disease are at high risk of central nervous system complications that can cause substantial morbidity. The condition's neurological effects are linked to a single amino acid substitution in the hemoglobin chain, and brain infarction can occur early in life. The supplied pediatric review further reports that hemorrhagic brain complications become more common as patients age.
Living With Neurological Risk
Neurological complications can affect people with sickle cell disease through both acute events and longer-term consequences. The human impact may extend beyond clinically obvious stroke to less visible problems that affect brain function and quality of life. The supplied material does not provide patient narratives or region-specific experience data, so the personal burden should be described cautiously.
The findings emphasize that neurological damage often begins early in life, underscoring the importance of early detection and intervention. Regular neuropsychological assessments and neuroimaging studies should be considered, even in asymptomatic patients, to identify silent infarcts and cognitive deficits.
While this review provides valuable insights, further research is needed to address the specific challenges of managing SCD in Africa. These challenges include limited resources, inadequate screening, and regional differences in healthcare standards. By addressing these gaps, we can improve the lives of those living with SCD and protect their neurological health.